

This is a nextflow process that predicts gene models in Caenorhabditis de novo assembled genomes using BRAKER3 guided with reference/Eukaryota proteomes.
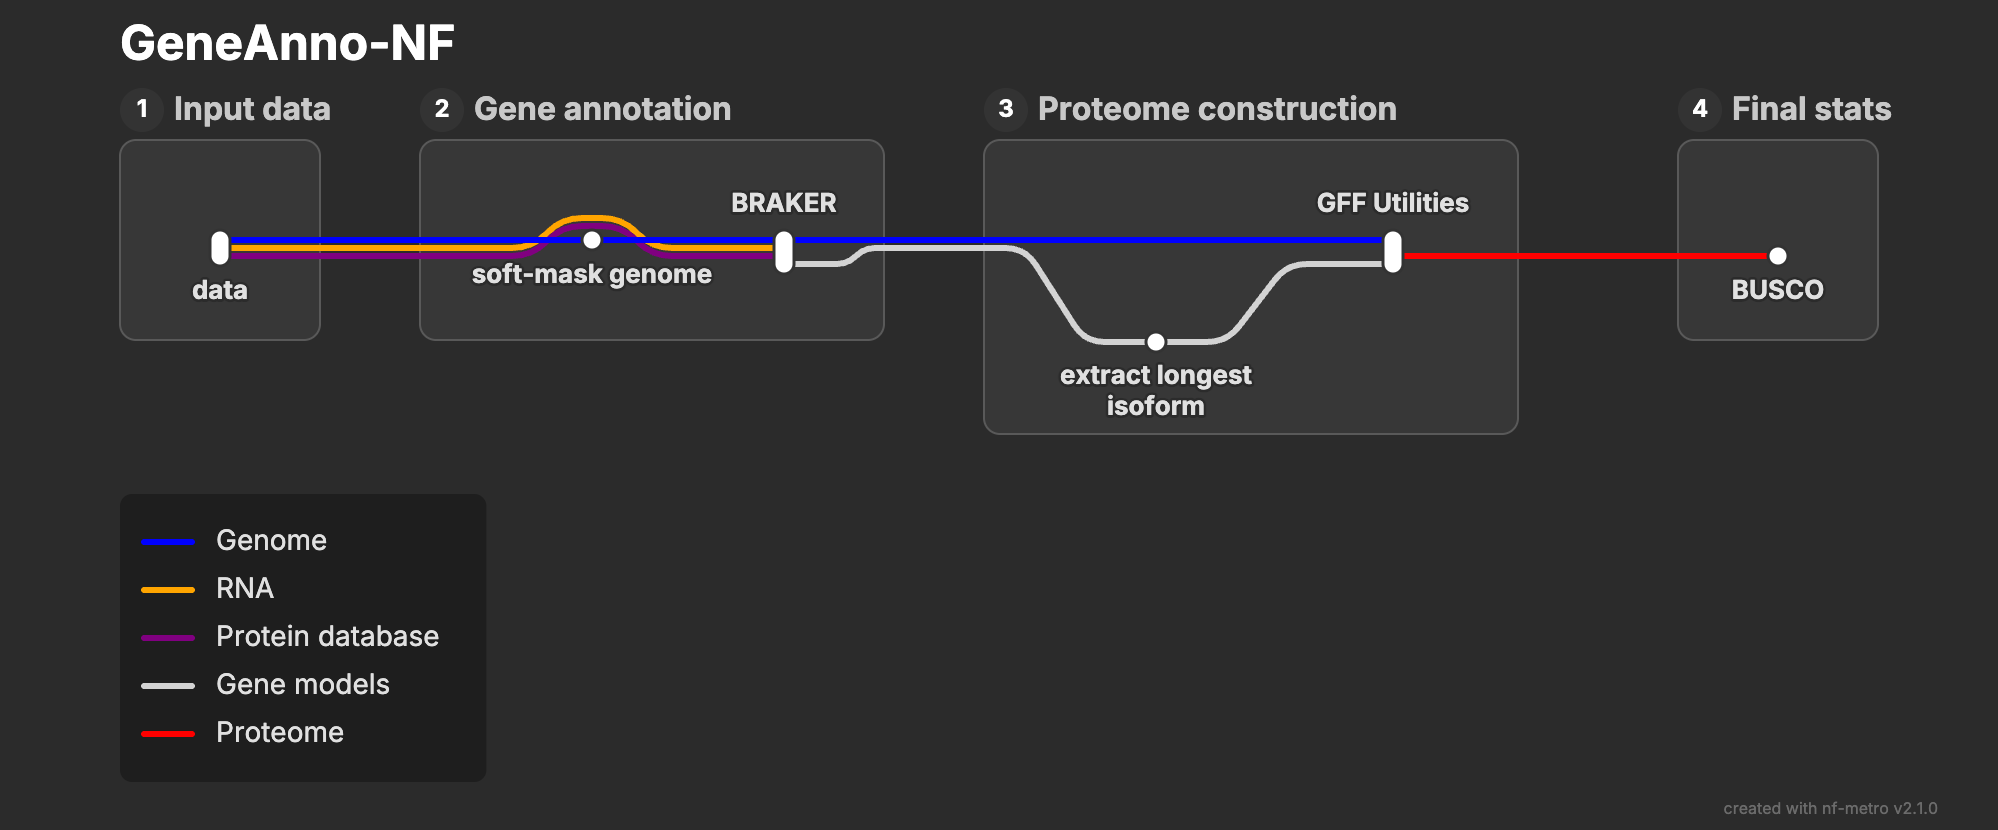
Workflow diagram generated using nf-metro.
This Nextflow process is meant to be run after assembly-nf, and it will add full paths to files produced and busco protein stats directly to the stats sheet output from assembly-nf.
The parameter --source can be equal to "only-braker" or "default". If --source is set to "only-braker", then only BRAKER3 will run. Otherwise, if --source is equal to "default", then the entire workflow will run, which is inclusive of BRAKER3, agat for extracting the longest isoform of each gene model prediction, gffread for creating a proteome from the longest isoform GFF and the supplied genome, BUSCO on the created proteome, and then gathering all paths and stats and adding to the supplied --sample_sheet.
The formatting of the supplied --sample_sheet is dependent on what --source is equal to. If --source is equal to "only-braker", then the provided --sample_sheet should be a TSV with headers: species, strain, asm_path. If --source is equal to "default", then the provided --sample_sheet can either be the stats TSV output from assembly-nf, or a custom TSV with any number of headers, but it must contain the required columns of: species, strain, asm_path. The absolute path to the --sample_sheet needs to be provided.
The parameter --species can be equal to "c_elegans", "c_tropicalis", "c_briggsae", or "c_nigoni". The --species specifies which proteome/protein database to use for guided gene model predictions. Additionally, the specified species will determine which file is used for soft-masking an assembly prior to running BRAKER3.
The parameter -profile specifies the configuration of the pipeline, inclusive of default parameters, resource allocation, and process-specific environments/containers. The only option for -profile right now is "rockfish". The user can created their own config file in /geneAnno-nf/conf/ if they wish to run on their HPC.
The parameters --mode is used to specify what sources of information you wish to have BRAKER3 use to guide gene model prediction. The user must set the mode to "prot", "rna_and_prot", or "rna" to indicate if BRAKER3 should use proteins to guide gene model prediction (previously benchmarked protein databases are used based on the --species parameter), short-read RNA-seq and proteins, or just short-read RNA-seq. If the user does not specify any --mode, then the pipeline will run in default mode with only proteins guiding gene model creation. If --mode "rna" or "rna_and_prot" are specified, the user must provide the full path to the BAM file containing RNA-alignment information in the --sample_sheet with column header "bam".
nextflow run main.nf --source default --sample_sheet $PWD/nigoni_asm-nf_filteredStats.tsv --outdir nigoni_geneModels --species c_nigoni --mode prot -profile rockfish